Resource
| Id | pipeline/GPF_SFARI_annotation |
|---|---|
| Type | annotation_pipeline |
| Version | 0 |
| Summary | GPF SFARI Annotation Pipeline |
| Description |
This is a pipeline to annotate with autism related attributes. |
| Labels |
Pipeline Documentation
preamble
| Summary | GPF SFARI Annotation Pipeline |
|---|---|
| Description | This is the pipeline used to annotate SFARI GPF instance resources |
| Input reference genome | hg38/genomes/GRCh38.p14 |
Annotators
Worst effect across all transcripts.
comma separated list of genes with worst effect.
<gene_1>:<effect_1>|... A gene can be repeated.
Effect details for each affected transcript. Format: < transcript 1 >:<gene 1>:<effect 1>:<details 1>|...
List of all genes
Annotator to identify the effect of the variant on protein coding.
Normalized allele.
dbSNP ID (i.e. rs number)
aggregator: list
Annotator to use with scores that depend on allele like variant frequencies, etc.
Mode (mode parameter, applies to VCFAllele inputs only):
allele(default): routed by the resource'sallele_score_mode. On anallelesresource every allele is an exact chrom/pos/ref/alt match. On asubstitutionsresource only a substitution is; an insertion, deletion or complex allele aggregates the allele lines over the bases it covers.region: aggregates scores for all allele lines overlapping the annotatable's span, whatever the resource's mode.
Non-VCFAllele annotatables always use region aggregation.
- input_annotatable:
normalized_allele
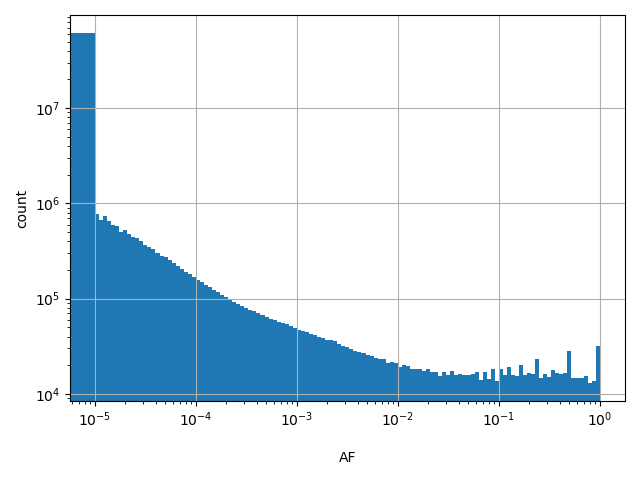
Alternate allele frequency
aggregator: max
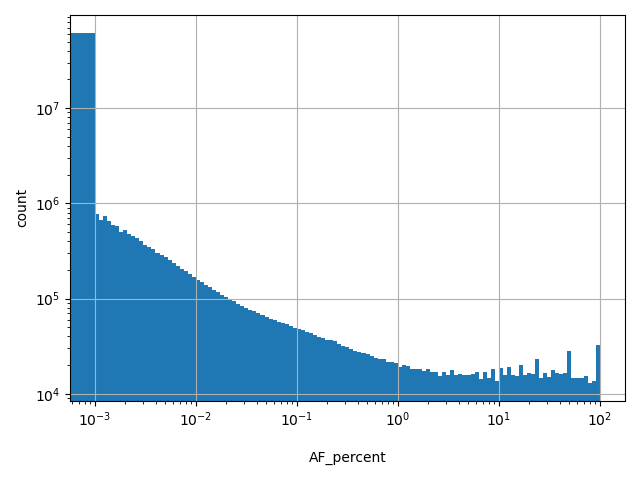
Alternate allele frequency as percent
aggregator: max
Annotator to use with scores that depend on allele like variant frequencies, etc.
Mode (mode parameter, applies to VCFAllele inputs only):
allele(default): routed by the resource'sallele_score_mode. On anallelesresource every allele is an exact chrom/pos/ref/alt match. On asubstitutionsresource only a substitution is; an insertion, deletion or complex allele aggregates the allele lines over the bases it covers.region: aggregates scores for all allele lines overlapping the annotatable's span, whatever the resource's mode.
Non-VCFAllele annotatables always use region aggregation.
- input_annotatable:
normalized_allele
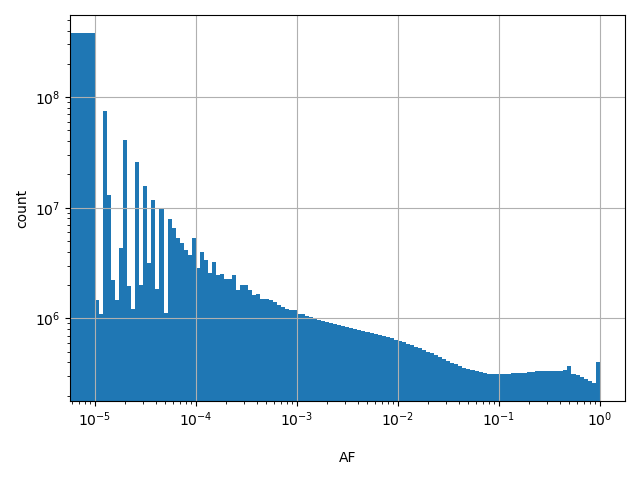
Alternate allele frequency
aggregator: max
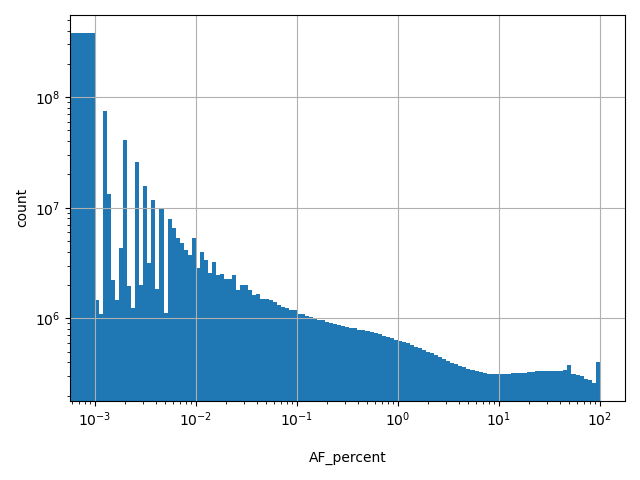
Alternate allele frequency as percent
aggregator: max
Annotator to use with scores that depend on allele like variant frequencies, etc.
Mode (mode parameter, applies to VCFAllele inputs only):
allele(default): routed by the resource'sallele_score_mode. On anallelesresource every allele is an exact chrom/pos/ref/alt match. On asubstitutionsresource only a substitution is; an insertion, deletion or complex allele aggregates the allele lines over the bases it covers.region: aggregates scores for all allele lines overlapping the annotatable's span, whatever the resource's mode.
Non-VCFAllele annotatables always use region aggregation.
- input_annotatable:
normalized_allele
ClinVar's preferred disease name for the concept specified by disease identifiers in CLNDISDB
aggregator: list
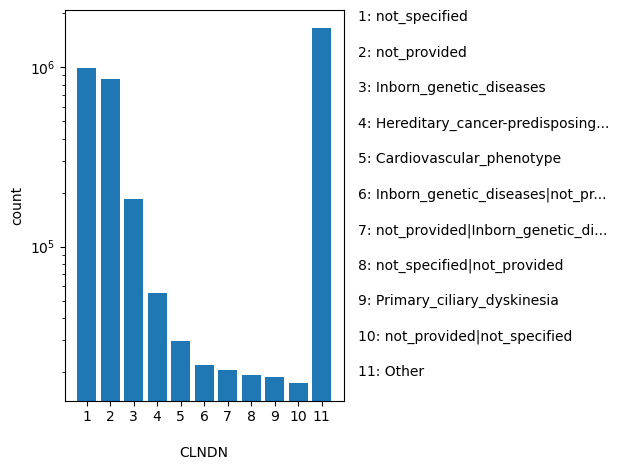
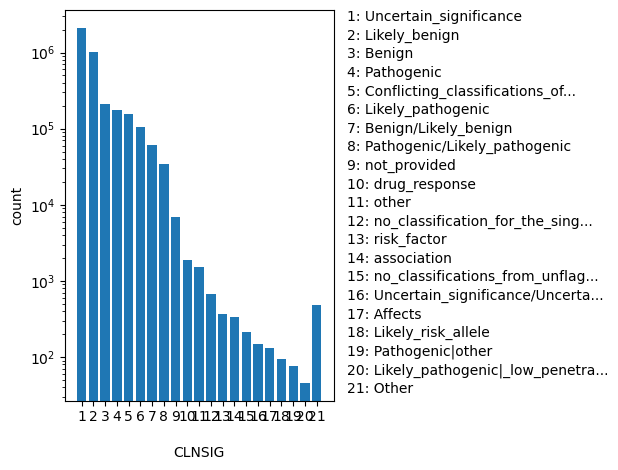
Aggregate germline classification for this single variant; multiple values are separated by a vertical bar
aggregator: list
Annotator to use with scores that depend on allele like variant frequencies, etc.
Mode (mode parameter, applies to VCFAllele inputs only):
allele(default): routed by the resource'sallele_score_mode. On anallelesresource every allele is an exact chrom/pos/ref/alt match. On asubstitutionsresource only a substitution is; an insertion, deletion or complex allele aggregates the allele lines over the bases it covers.region: aggregates scores for all allele lines overlapping the annotatable's span, whatever the resource's mode.
Non-VCFAllele annotatables always use region aggregation.
- input_annotatable:
normalized_allele
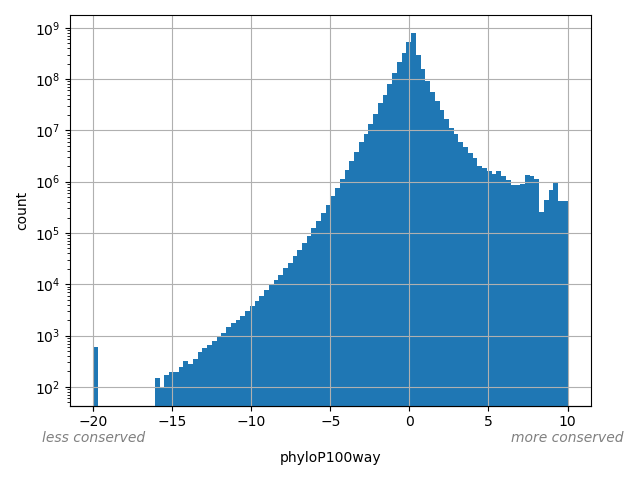
The score is a number that reflects the conservation at a position.
aggregator: mean [default]
Annotator to use with genomic scores depending on genomic position like phastCons, phyloP, FitCons2, etc.
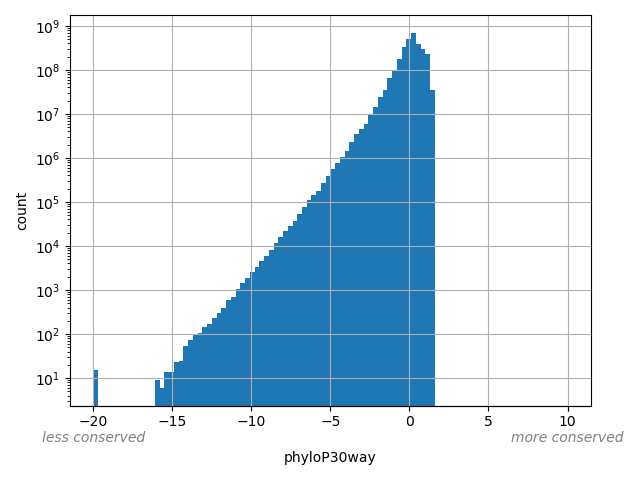
The score is a number that reflects the conservation at a position.
aggregator: mean [default]
Annotator to use with genomic scores depending on genomic position like phastCons, phyloP, FitCons2, etc.
The score is a number that reflects the conservation at a position.
aggregator: mean [default]
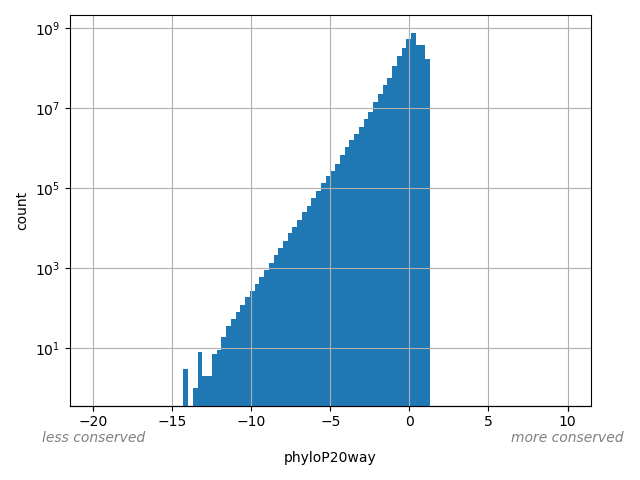
Annotator to use with genomic scores depending on genomic position like phastCons, phyloP, FitCons2, etc.
The score is a number that reflects the conservation at a position.
aggregator: mean [default]
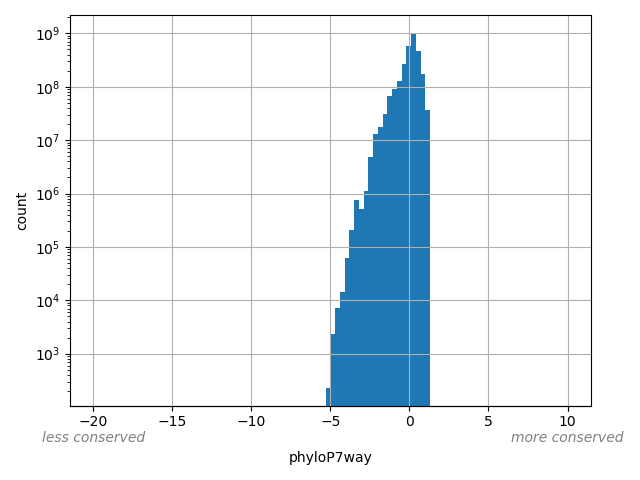
Annotator to use with genomic scores depending on genomic position like phastCons, phyloP, FitCons2, etc.
The score is a number that reflects the conservation at a position.
aggregator: mean [default]
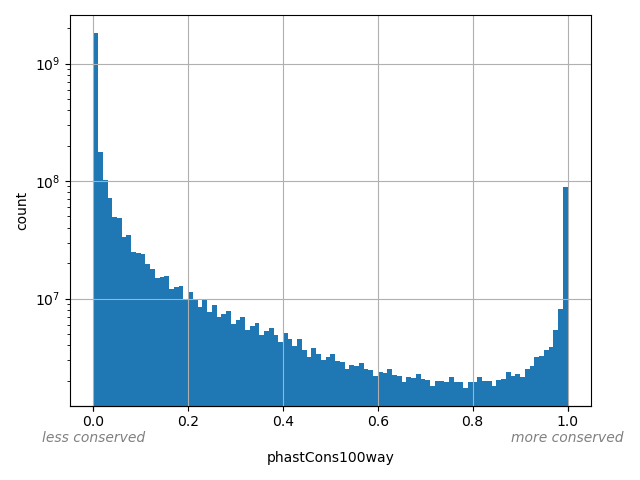
Annotator to use with genomic scores depending on genomic position like phastCons, phyloP, FitCons2, etc.
The score is a number that reflects the conservation at a position.
aggregator: mean [default]
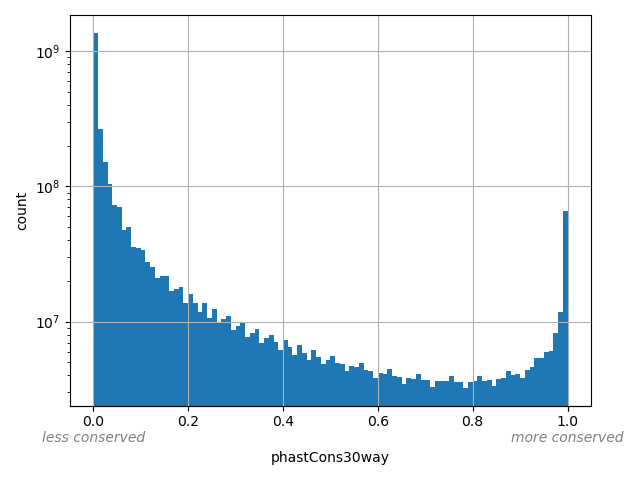
Annotator to use with genomic scores depending on genomic position like phastCons, phyloP, FitCons2, etc.
The score is a number that reflects the conservation at a position.
aggregator: mean [default]
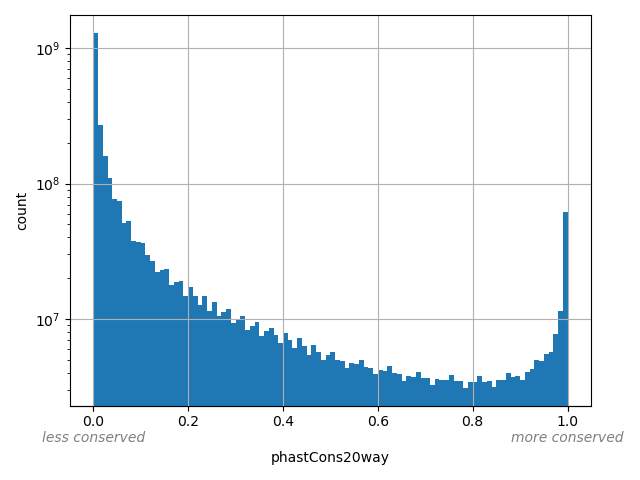
Annotator to use with genomic scores depending on genomic position like phastCons, phyloP, FitCons2, etc.
The score is a number that reflects the conservation at a position.
aggregator: mean [default]
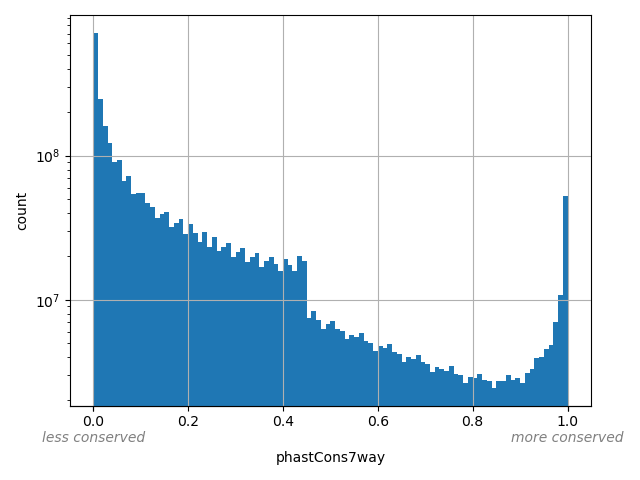
Annotator to use with genomic scores depending on genomic position like phastCons, phyloP, FitCons2, etc.
CADD raw score for functional prediction of a SNP. The larger the score the more likely the SNP has damaging effect
aggregator: max
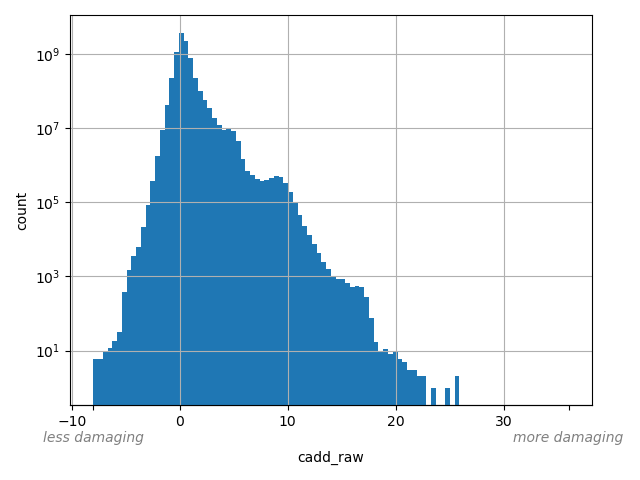
CADD phred-like score. This is phred-like rank score based on whole genome CADD raw scores. The larger the score the more likely the SNP has damaging effect.
aggregator: max
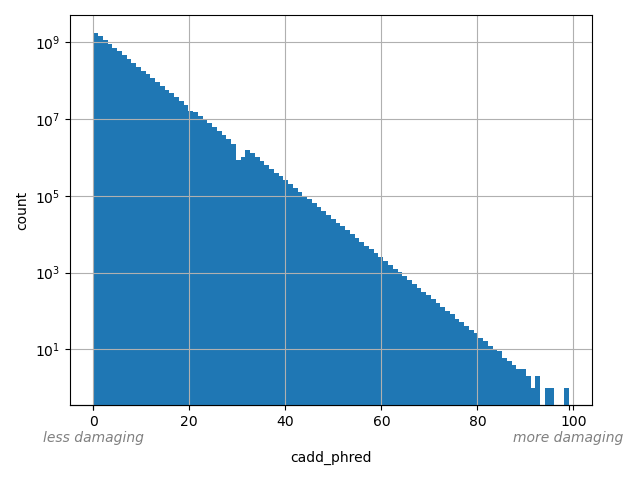
Annotator to use with scores that depend on allele like variant frequencies, etc.
Mode (mode parameter, applies to VCFAllele inputs only):
allele(default): routed by the resource'sallele_score_mode. On anallelesresource every allele is an exact chrom/pos/ref/alt match. On asubstitutionsresource only a substitution is; an insertion, deletion or complex allele aggregates the allele lines over the bases it covers.region: aggregates scores for all allele lines overlapping the annotatable's span, whatever the resource's mode.
Non-VCFAllele annotatables always use region aggregation.
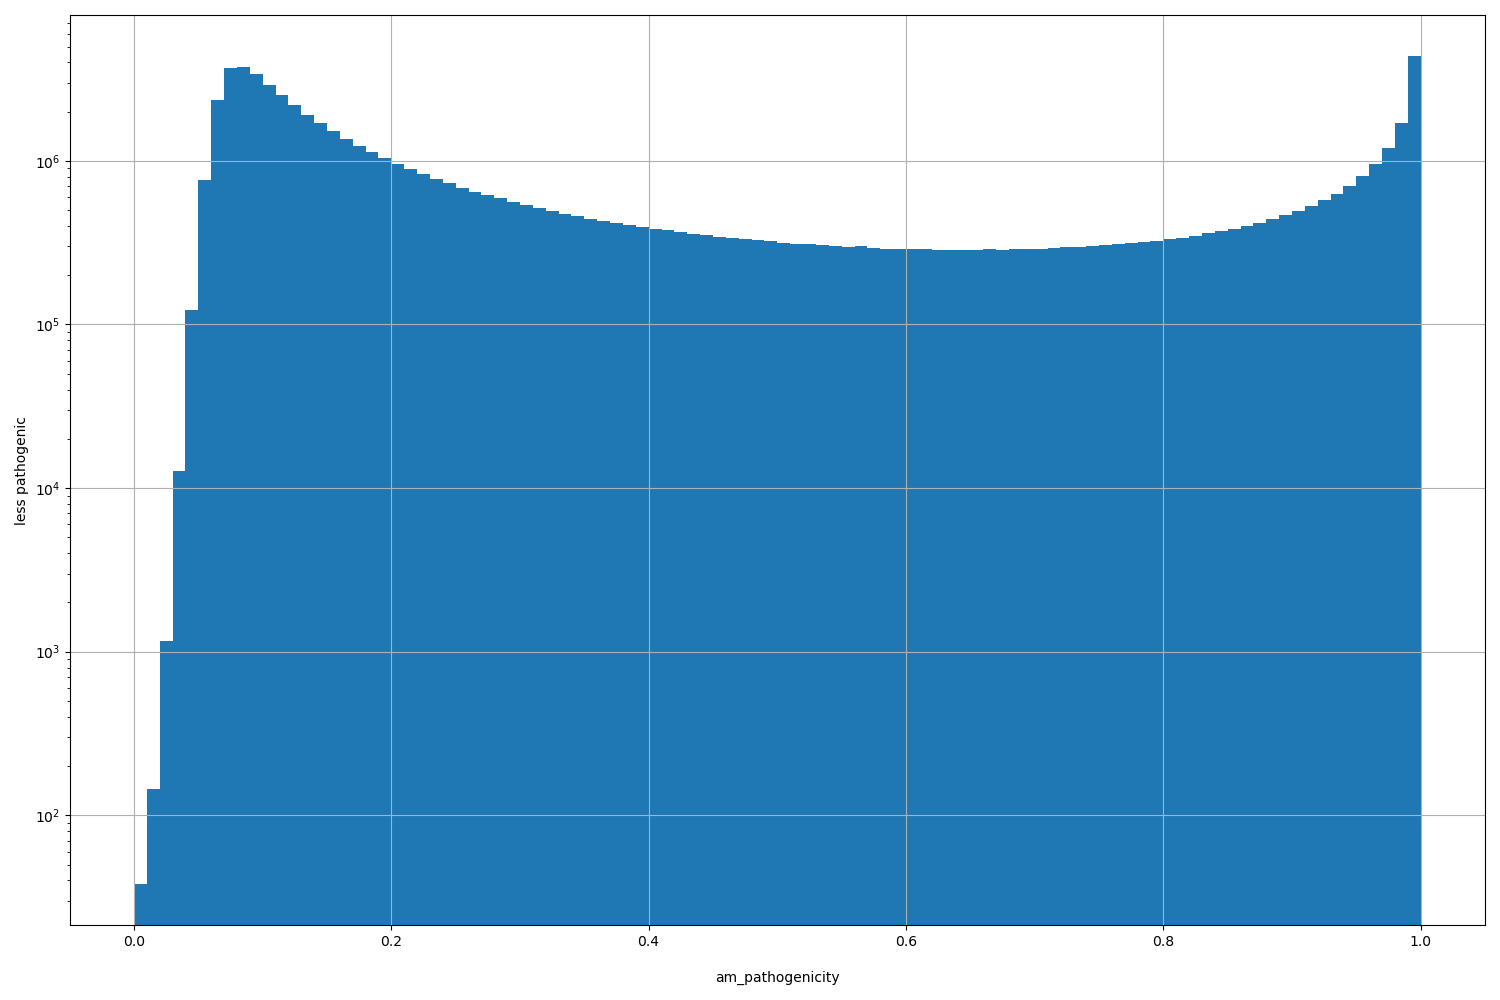
AlphaMissense Pathogenicity score is a deleteriousness score for missense variants
aggregator: max
Annotator to use with scores that depend on allele like variant frequencies, etc.
Mode (mode parameter, applies to VCFAllele inputs only):
allele(default): routed by the resource'sallele_score_mode. On anallelesresource every allele is an exact chrom/pos/ref/alt match. On asubstitutionsresource only a substitution is; an insertion, deletion or complex allele aggregates the allele lines over the bases it covers.region: aggregates scores for all allele lines overlapping the annotatable's span, whatever the resource's mode.
Non-VCFAllele annotatables always use region aggregation.
The lifted over annotatable
Annotator to lift over a variant from one reference genome to another.
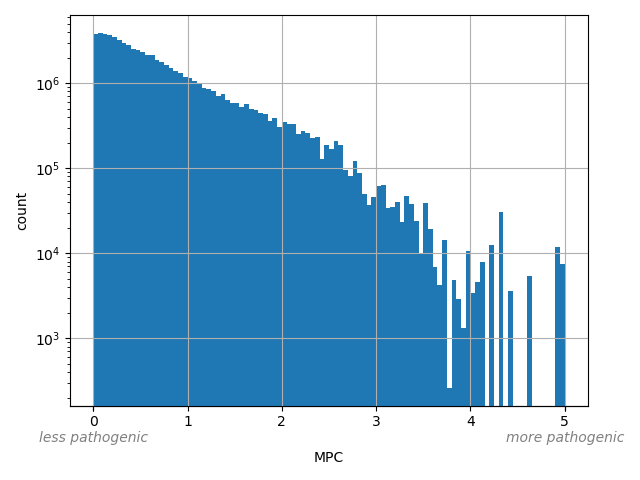
Missense badness, PolyPhen-2, and Constraint. A deleteriousness prediction score for missense variants"
aggregator: max
Annotator to use with scores that depend on allele like variant frequencies, etc.
Mode (mode parameter, applies to VCFAllele inputs only):
allele(default): routed by the resource'sallele_score_mode. On anallelesresource every allele is an exact chrom/pos/ref/alt match. On asubstitutionsresource only a substitution is; an insertion, deletion or complex allele aggregates the allele lines over the bases it covers.region: aggregates scores for all allele lines overlapping the annotatable's span, whatever the resource's mode.
Non-VCFAllele annotatables always use region aggregation.
- input_annotatable:
hg19_annotatable
Worst effect across all transcripts.
Effect details for each affected transcript. Format: < transcript 1 >:<gene 1>:<effect 1>:<details 1>|...
<gene_1>:<effect_1>|... A gene can be repeated.
Annotator to identify the effect of the variant on protein coding.
The number of fragments overlapping the annotatable and kept by the fragment filter.
The number of fragments overlapping the annotatable and kept by the fragment filter.
The number of fragments overlapping the annotatable and kept by the fragment filter.
Files
| Filename | Size | md5 |
|---|---|---|
| annotation.yaml | 3.33 KB | d33cf4e342bd7596bd5c0fe812f46741 |
| genomic_resource.yaml | 196.0 B | d0b001f30fff2e7ed506b17cdfb008ed |
| statistics/ |